This study links an epigenetic mechanism in macrophages (H2A.Z deposition) to microbial metabolites (butyrate) and to genetic variants associated with IBD, which may help explain persistent inflammation and variable treatment responses.
Who Should Pay Attention
Researchers in IBD immunology and microbiome science; clinicians interested in mechanisms of anti-TNF nonresponse; patients curious about research into IBD causes.
Study Snapshot
Story typeResearch paper
Evidence typeResearch paper
Source depthJournal abstract
What To Know
This study (abstract provided) describes a mechanism in macrophages where the histone variant H2A.Z is removed during activation and re-deposited during resolution; mice lacking myeloid H2A.Z showed excessive inflammation, worse colitis, and loss of butyrate-producing gut bacteria.
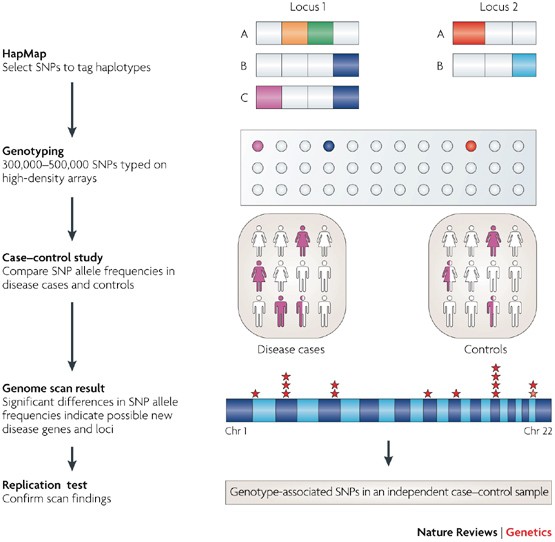
The authors link human IBD risk variants to lower expression of GAS41 and TIP60, a reader-writer module needed for H2A.Z deposition, and note that lower GAS41/TIP60 in patient tissues correlates with disease progression and poor anti-TNF response.
Keep In Mind
Findings combine mouse genetic models, mechanistic cell biology, microbiome changes, and correlative human tissue expression data; as basic research, it suggests pathways for future translational work but does not establish clinical treatments.
Source Details
Review the original publication for the complete reporting, methods, and context.
Research paperEvidence type derived from source or registry metadata.
PublicationProtein & cell
AuthorsZhaoran Sun, Fanyi Meng, Chunjian Piao +9 more
InstitutionState Key Laboratory of Experimental Hematology, The Province and Ministry Co-sponsored Collaborative Innovation Center for Medical Epigenetics, Tianjin Key Laboratory of Medical Epigenetics, Key Laboratory of Immune Microenvironment and Disease (Ministry of Education), Department of Cell Biology, School of Basic Medical Sciences, Tianjin Medical University, Tianjin 300070, China.
This Cure8 brief is based on source text from the linked article. Cure8 is informational only and is not a substitute for professional medical advice, diagnosis, or treatment.
The work links STAT4 — a gene locus associated with IBD — to Th17 pathogenic behavior and suppression of the anti-inflammatory cytokine IL-10, offering a possible mechanism by which STAT4 variants could worsen gut...
IBD patients face higher risk of osteoporosis; identifying shared, apoptosis-related genes may help explain why and point to future biomarkers or therapeutic targets. The study links specific genes (WNT1, S100A8) to both...
Genetic studies help reveal biological pathways behind Crohn’s disease, which can guide research into new treatments and better understanding of disease mechanisms. People with Crohn’s disease may find it useful to know...